AgP2
AgP2 is a stable semiconducting compound composed of silver and phosphorus.
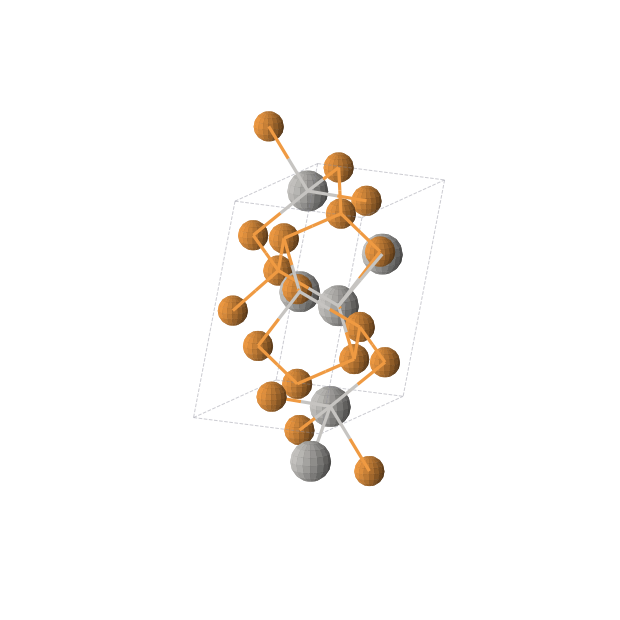
About AgP2
AgP2 is a silver phosphide compound that exhibits semiconducting electronic properties. As a thermodynamically stable material situated on the convex hull, it represents a robust phase within the silver-phosphorus system.
Its structural complexity is highlighted by its presence across numerous reported experimental and computational databases. This widespread documentation underscores its significance as a subject of investigation for researchers exploring functional inorganic semiconductors.
Key Properties
Cross-validated computational properties for AgP2, aggregated across 7 databases.
Band GapEnergy needed to move an electron from the valence band to the conduction band. Lower or zero values tend to behave more metallic; larger gaps are more insulating or semiconducting.
Energy Above HullThermodynamic distance from the most stable set of competing phases. 0 eV/atom is on the convex hull; small positive values may still be experimentally accessible.
StabilityA plain-language summary of the best reported energy-above-hull result. It reflects whether the lowest-energy structure is on, near, or far from the stability hull.
StructuresCount of reported calculated crystal structures for this formula, including alternate polymorphs, source databases, and observed space groups.
Cross-Source DFT Agreement
How well independent DFT databases agree on the thermodynamics of AgP2. Tight agreement means computed properties can be trusted without re-running calculations.
Agreement ScoreA normalized confidence score summarizing how closely independent DFT databases agree. Higher scores mean tighter cross-source agreement.
Hull SpreadDifference between the highest and lowest energy-above-hull values reported by comparable sources. Smaller spread means less thermodynamic disagreement.
Sources ComparedNumber and names of computational sources with comparable entries for this formula.
Space Group ConsensusWhether independent sources predict the same crystal symmetry for the lowest-energy structure.
Reported Structures
Lowest-energy structures reported for AgP2, ranked by energy above hull.
| Space GroupSymmetry classification of the crystal arrangement. The number is the international space-group index. | Crystal SystemBroad lattice family, such as cubic, tetragonal, monoclinic, or triclinic, derived from unit-cell symmetry. | Band Gap (eV)Electronic gap calculated for this specific reported structure, measured in electronvolts. | E above hull (eV/atom)Thermodynamic distance from the convex hull for this structure, normalized per atom. Lower is generally more stable. | E/atom (eV)Computed total energy normalized per atom. Use energy above hull, not this value alone, when comparing stability. | Density (g/cm³)Mass per relaxed crystal volume, reported in grams per cubic centimeter. |
|---|---|---|---|---|---|
| P21/c (No. 14) | monoclinic | 0.66 | 0.0009 | -13.158 | 4.98 |
| P21/c (No. 14) | monoclinic | 0.67 | 0.0119 | -0.005 | — |
| — | — | 0.68 | 0.0133 | -0.178 | — |
| P-1 (No. 2) | triclinic | 0.00 | 0.1823 | 0.165 | — |
| Pmmm (No. 47) | orthorhombic | 0.00 | 0.2855 | 0.268 | — |
| — | — | 0.00 | 0.6323 | 0.441 | — |
| — | — | 0.00 | 1.7657 | 1.575 | — |
| C2/m (No. 12) | monoclinic | — | — | — | 5.84 |
| — | — | — | — | — | 4.83 |
| C2/m (No. 12) | monoclinic | — | — | — | 4.46 |
| P-1 (No. 2) | triclinic | — | — | — | 7.74 |
| P-1 (No. 2) | triclinic | — | — | — | 3.46 |
Applications
Where AgP2 is used.
Frequently Asked Questions
Common questions about AgP2, answered from cross-validated data.
What is AgP2?
AgP2 is a stable semiconducting compound composed of silver and phosphorus.
What is AgP2 used for?
What is the band gap of AgP2?
Is AgP2 a metal, semiconductor, or insulator?
Is AgP2 thermodynamically stable?
What is the crystal structure of AgP2?
What is the density of AgP2?
How many polymorphs of AgP2 are known?
What elements does AgP2 contain?
Where does the data for AgP2 come from?
How It Compares
As a distinct silver phosphide, AgP2 serves as a foundational reference point for understanding the binary chemistry of silver and phosphorus. Without direct structural siblings in this specific class, it stands as a unique example of how these elements combine into a stable, semiconducting framework.
Data sources & attribution
- materials_project — Data from the Materials Project (materialsproject.org). Cite: Jain et al., APL Materials 1, 011002 (2013). (CC-BY-4.0)
- jarvis — Data from JARVIS (jarvis.nist.gov), NIST. Cite: Choudhary et al., npj Comp. Mater. 6, 173 (2020). (LicenseRef-US-Gov-PD)
- oqmd — Data from the OQMD (oqmd.org). Cite: Saal et al., JOM 65, 1501 (2013); Kirklin et al., npj Comp. Mater. 1, 15010 (2015). (CC-BY-4.0)
- mpaloe — Data from MP-ALOE. Cite: Kuner et al., npj Comput. Mater. (2025), doi:10.1038/s41524-025-01834-9.
- omat24 — Data from Meta Open Materials 2024 (OMat24), Meta FAIR. Cite: Barroso-Luque et al., arXiv:2410.12771 (2024). (CC-BY-4.0)
Analyze AgP2 in the Lattice Graph platform
Polymorph comparison, confidence scoring, supply-chain risk, and patent monitoring — across 85 source databases.
Explore the Platform →